Systematics - Phylogeny of invertebrates based on genomic data (InvertOmics)
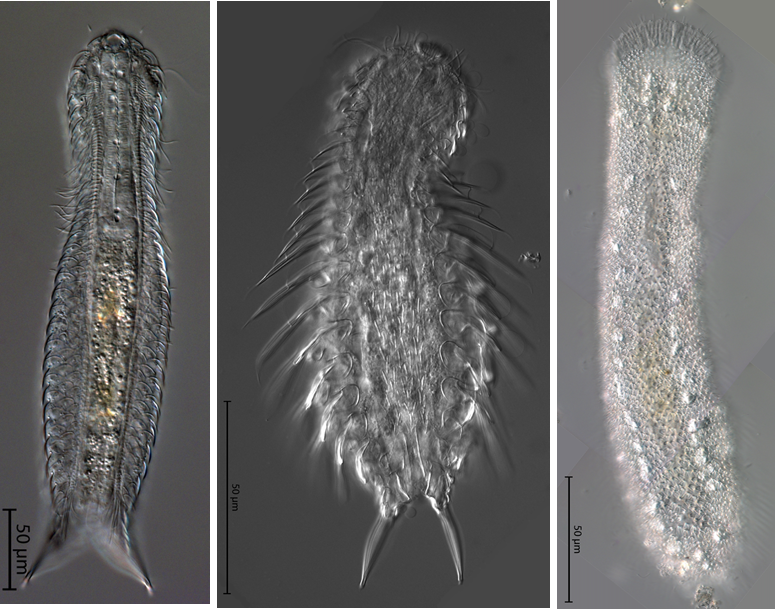
How the last common ancestor of animals like humans, fishes, insects, roundworms, snails and earthworms looked and how evolution proceeded within this large group of animals is a major knowledge gap in biology. To resolve this gap, this project seeks to understand how the phyla in the superphylum Lophotrochozoa, comprising 16 of the 32 animal phyla, are related to each other as this is presently unknown.
Torsten Struck, UiO
About the project
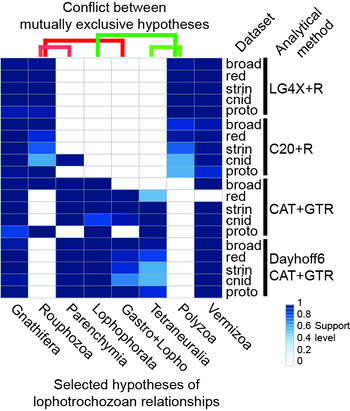
The origin and evolution of Bilateria including, among others, humans, fishes, insects, roundworms, snails and earthworms is controversially discussed in several biological disciplines such as systematics or evolutionary developmental biology. In one hypothesis, evolution in Bilateria advances from a simple body organization similar to flatworms towards more complex forms several times independently. In the other one, the evolution progresses in the opposite direction from a complex ancestor more like an annelid to simple organizations by several separate reductions. Support for one or the other depends on the phylogeny and evolution of Lophotrochozoa, one of the major bilaterian taxa, but a robust phylogeny is still lacking despite recent phylogenomic studies. This is due to both low coverage by genomic data and misleading biases in data of lophotrochozoan taxa.
In this project, high-quality reference genomes shall be generated for most lophotrochozoan phyla using high-throughput sequencing. As several lophotrochozoan species have very small body sizes (< 500 μm length) this is challenging, but we are tackling this by implementing protocols from single-cell genomics developed for other disciplines like cancer research. This means that genomic data derived from single small-sized individuals be used for phylogenomic studies for the first time in Lophotrochozoa. With this, we will also contribute to the goal of the Earth Biogenome Project (EBP) (external link) to sequence a reference genome of each single eukrayotic species on Earth.
Another challenge in the reconstruction of the phylogeny of Lophotrochozoa are different biases in the data that may mislead the analysis. Using simulated and empirical data, we will develop tools tailored towards such problems and establish both best procedures ameliorating such negative effects of biases and a new tools to detect such biases in the data in an efficient way correcting of possible influence such as the kind of data used (nucleotide vs. amino acid), and the number of taxa and positions analyzed.
Due to both the large genomic dataset and these thorough analyses, a robust phylogeny of Lophotrochozoa will be provided allowing contributions to discussions about the origin and evolution of Bilateria as well as of lophotrochozoan taxa and character traits. The project will generate deep genomic resources for several non-model organisms providing, e.g., new opportunities for studies in functional genome evolution, developmental biology, ecology or discovery of new medicinal drugs.
Roberts, Nickellaus G; Tweedt, Sarah M; Meyer, Christopher; Struck, Torsten H & Kocot, Kevin M
(2025).
Chromosome-level haplotype-resolved genome assembly of the linguliform brachiopod Discradisca antillarum (d'Orbigny, 1845).
G3: Genes, Genomes, Genetics.
15(12).
doi: 10.1093/g3journal/jkaf233.
Full text in Research ArchiveShow summary
Brachiopods are a group of shelled, filter-feeding marine invertebrates. Though they superficially resemble bivalve mollusks, brachiopods fall within the clade Lophophorata (Brachiopoda, Bryozoa, Phoronida) and possess an abundant and diverse fossil record spanning more than 500 million years. This long evolutionary history makes Brachiopoda particularly important for understanding the evolution of the morphologically disparate superphylum Lophotrochozoa. Brachiopods also stand to provide insight into the evolution of biomineralization, as most lineages produce skeletons of calcium carbonate while linguliforms use calcium phosphate. However, genomic resources for brachiopods are limited. To enable comparative and evolutionary genomic studies of this clade, we generated a chromosome-level assembly for the discinid brachiopod Discradisca antillarum. Sequencing was performed using PacBio HiFi and Hi-C to produce both a primary (N = 8) and haplotype-resolved (2N = 16) assembly. The primary assembly comprises 208 scaffolds with the 8 chromosome-level scaffolds representing 96.6% of the genome. The primary assembly has a BUSCO genome completeness score of 95.5% (94.4% single copy, 1.0% duplicated, 0.9% fragmented). Annotation of protein-coding genes yielded 29,208 genes with a BUSCO protein completeness score of 95.6% (87.6% single copy, 8.0% duplicated, 2.8% fragmented). Comparative synteny between D. antillarum and other lophotrochozoans shows that, at the macrosyntenic level, the genome conserves all proposed ancestral lophotrochozoan fusion-with-mixing events while also revealing new fusions involving several bilaterian ancestral linkage groups. The genome of D. antillarum will enable significant insights into brachiopod evolution.
Gracia, Alberto Valero; Roberts, Nickellaus G; Yap-Chiongco, Meghan; Capucho, Ana Teresa; Kocot, Kevin M & Matschiner, Michael
[Show all 7 contributors for this article](2024).
First Chromosome-Level Genome Assembly of a Ribbon Worm from the Hoplonemertea Clade, Emplectonema gracile, and Its Structural Annotation.
Genome Biology and Evolution (GBE).
16(7).
doi: 10.1093/gbe/evae127.
Full text in Research ArchiveShow summary
Genome-wide information has so far been unavailable for ribbon worms of the clade Hoplonemertea, the most species-rich class within the phylum Nemertea. While species within Pilidiophora, the sister clade of Hoplonemertea, possess a pilidium larval stage and lack stylets on their proboscis, Hoplonemertea species have a planuliform larva and are armed with stylets employed for the injection of toxins into their prey. To further compare these developmental, physiological, and behavioral differences from a genomic perspective, the availability of a reference genome for a Hoplonemertea species is crucial. Such data will be highly useful for future investigations toward a better understanding of molecular ecology, venom evolution, and regeneration not only in Nemertea but also in other marine invertebrate phyla. To this end, we herein present the annotated chromosome-level genome assembly for Emplectonema gracile (Nemertea; Hoplonemertea; Monostilifera; Emplectonematidae), an easily collected nemertean well suited for laboratory experimentation. The genome has an assembly size of 157.9 Mb. Hi-C scaffolding yielded chromosome-level scaffolds, with a scaffold N50 of 10.0 Mb and a score of 95.1% for complete BUSCO genes found as a single copy. Annotation predicted 20,684 protein-coding genes. The high-quality reference genome reaches an Earth BioGenome standard level of 7.C.Q50.
A genomic database of all Earth’s eukaryotic species could contribute to many scientific discoveries; however, only a tiny fraction of species have genomic information available. In 2018, scientists across the world united under the Earth BioGenome Project (EBP), aiming to produce a database of high-quality reference genomes containing all ~1.5 million recognized eukaryotic species. As the European node of the EBP, the European Reference Genome Atlas (ERGA) sought to implement a new decentralised, equitable and inclusive model for producing reference genomes. For this, ERGA launched a Pilot Project establishing the first distributed reference genome production infrastructure and testing it on 98 eukaryotic species from 33 European countries. Here we outline the infrastructure and explore its effectiveness for scaling high-quality reference genome production, whilst considering equity and inclusion. The outcomes and lessons learned provide a solid foundation for ERGA while offering key learnings to other transnational, national genomic resource projects and the EBP.
Roberts, Nickellaus G.; Gilmore, Michael J.; Struck, Torsten H & Kocot, Kevin M.
(2024).
Multiple Displacement Amplification Facilitates SMRT Sequencing of Microscopic Animals and the Genome of the Gastrotrich Lepidodermella squamata (Dujardin 1841).
Genome Biology and Evolution (GBE).
16(12).
doi: 10.1093/gbe/evae254.
Full text in Research ArchiveShow summary
Obtaining adequate DNA for long-read genome sequencing remains a roadblock to producing contiguous genomes from small-bodied organisms, hindering understanding of phylogenetic relationships and genome evolution. Multiple displacement amplification leverages Phi29 DNA polymerase to produce micrograms of DNA from picograms of input. However, multiple displacement amplification's inherent biases in amplification related to guanine and cytosine (GC) content, repeat content and chimera production are a problem for long-read genome assembly, which has been little investigated. We explored the utility of multiple displacement amplification for generating template DNA for High Fidelity (HiFi) sequencing directly from living cells of Caenorhabditis elegans (Nematoda) and Lepidodermella squamata (Gastrotricha) containing one order of magnitude less DNA than required for the PacBio Ultra-Low DNA Input Workflow. High Fidelity sequencing of libraries prepared from multiple displacement amplification products resulted in highly contiguous and complete genomes for both C. elegans (102 Mbp assembly; 336 contigs; N50 = 868 kbp; L50 = 39; BUSCO_nematoda_nucleotide: S:96.1%, D:2.8%) and L. squamata (122 Mbp assembly; 157 contigs; N50 = 3.9 Mbp; L50 = 13; BUSCO_metazoa_nucleotide: S:80.8%, D:2.8%). Coverage uniformity for reads from multiple displacement amplification DNA (Gini Index: 0.14, normalized mean across all 100 kbp blocks: 0.49) and reads from pooled nematode DNA (Gini Index: 0.16, normalized mean across all 100 kbp blocks: 0.49) proved similar. Using this approach, we sequenced the genome of the microscopic invertebrate L. squamata (Gastrotricha), the first of its phylum. Using the newly sequenced genome, we infer Gastrotricha's long-debated phylogenetic position as the sister taxon of Platyhelminthes and conduct a comparative analysis of the Hox cluster.
The European Reference Genome Atlas (ERGA) consortium aims to generate a reference genome catalogue for all of Europe's eukaryotic biodiversity. The biological material underlying this mission, the specimens and their derived samples, are provided through ERGA’s pan-European network. To demonstrate the community’s capability and capacity to realise ERGA’s ambitious mission, the ERGA Pilot project was initiated. In support of the ERGA Pilot effort to generate reference genomes for European biodiversity, the ERGA Sampling and Sample Processing committee (SSP) was formed by volunteer experts from ERGA’s member base. SSP aims to aid participating researchers through (i) establishing standards for and collecting of sample/specimen metadata; (ii) prioritisation of species for genome sequencing; and (iii) development of taxon-specific collection guidelines including logistics support. SSP serves as the entry point for sample providers to the ERGA genomic resource production infrastructure and guarantees that ERGA’s high-quality standards are upheld throughout sample collection and processing. With the volume of researchers, projects, consortia, and organisations with interests in genomics resources expanding, this manuscript shares important experiences and lessons learned during the development of standardised operational procedures and sample provider support. The manuscript details our experiences in incorporating the FAIR and CARE principles, species prioritisation, and workflow development, which could be useful to individuals as well as other initiatives.
Fleming, James Frederick & Struck, Torsten H
(2023).
nRCFV: a new, dataset-size-independent metric to quantify compositional heterogeneity in nucleotide and amino acid datasets.
BMC Bioinformatics.
24.
doi: 10.1186/s12859-023-05270-8.
Full text in Research ArchiveShow summary
Motivation Compositional heterogeneity—when the proportions of nucleotides and amino acids are not broadly similar across the dataset—is a cause of a great number of phylogenetic artefacts. Whilst a variety of methods can identify it post-hoc, few metrics exist to quantify compositional heterogeneity prior to the computationally intensive task of phylogenetic tree reconstruction. Here we assess the efficacy of one such existing, widely used, metric: Relative Composition Frequency Variability (RCFV), using both real and simulated data. Results Our results show that RCFV can be biased by sequence length, the number of taxa, and the number of possible character states within the dataset. However, we also find that missing data does not appear to have an appreciable effect on RCFV. We discuss the theory behind this, the consequences of this for the future of the usage of the RCFV value and propose a new metric, nRCFV, which accounts for these biases. Alongside this, we present a new software that calculates both RCFV and nRCFV, called nRCFV_Reader. Availability and implementation nRCFV has been implemented in RCFV_Reader, available at: https://github.com/JFFleming/RCFV_Reader. Both our simulation and real data are available at Datadryad: https://doi.org/10.5061/dryad.wpzgmsbpn
Fleming, James Frederick
(2023).
The wealth of shared resources: Improving molecular taxonomy using eDNA and public databases.
Zoologica Scripta.
ISSN 0300-3256.
doi: 10.1111/zsc.12591.
Full text in Research Archive
Bachmann, Lutz; Beermann, Jan; Brey, Thomas; Boer, Hugo de; Dannheim, Jennifer & Edvardsen, Bente
[Show all 20 contributors for this article](2023).
The role of systematics for understanding ecosystem functions: Proceedings of the Zoologica Scripta Symposium, Oslo, Norway, 25 August 2022.
Zoologica Scripta.
ISSN 0300-3256.
52(3),
p. 187–214.
doi: 10.1111/zsc.12593.
Full text in Research ArchiveShow summary
Abstract On 25 August 2022, the Zoologica Scripta ‐ An International Journal of Systematic Zoology and the Norwegian Academy of Sciences and Letters arranged a symposium entitled ‘The role of systematics for understanding ecosystem functions’ in the Academy's premises in Oslo, Norway. The symposium aimed at offering a forum for exploring and discussing trends and future developments in the field of systematics. Eleven international experts contributed expertise on various issues related to global challenges, such as biodiversity assessments, databases, cutting‐edge analysis tools, and the consequences of the taxonomic impediment. Here, we compiled a multi‐author proceedings paper of the symposium contributions that are arranged in chapters and presents the content and the key conclusions of the majority of the presentations.
Struck, Torsten H; Golombek, Anja; Hoesel, Christoph; Dimitrov, Dimitar & Elgetany, Asmaa Haris
(2023).
Mitochondrial Genome Evolution in Annelida—A Systematic Study on Conservative and Variable Gene Orders and the Factors Influencing its Evolution.
Systematic Biology.
ISSN 1063-5157.
72(4),
p. 925–945.
doi: 10.1093/sysbio/syad023.
Full text in Research ArchiveShow summary
The mitochondrial genomes of Bilateria are relatively conserved in their protein-coding, rRNA, and tRNA gene complement, but the order of these genes can range from very conserved to very variable depending on the taxon. The supposedly conserved gene order of Annelida has been used to support the placement of some taxa within Annelida. Recently, authors have cast doubts on the conserved nature of the annelid gene order. Various factors may influence gene order variability including, among others, increased substitution rates, base composition differences, structure of noncoding regions, parasitism, living in extreme habitats, short generation times, and biomineralization. However, these analyses were neither done systematically nor based on well-established reference trees. Several focused on only a few of these factors and biological factors were usually explored ad-hoc without rigorous testing or correlation analyses. Herein, we investigated the variability and evolution of the annelid gene order and the factors that potentially influenced its evolution, using a comprehensive and systematic approach. The analyses were based on 170 genomes, including 33 previously unrepresented species. Our analyses included 706 different molecular properties, 20 life-history and ecological traits, and a reference tree corresponding to recent improvements concerning the annelid tree. The results showed that the gene order with and without tRNAs is generally conserved. However, individual taxa exhibit higher degrees of variability. None of the analyzed life-history and ecological traits explained the observed variability across mitochondrial gene orders. In contrast, the combination and interaction of the best-predicting factors for substitution rate and base composition explained up to 30% of the observed variability. Accordingly, correlation analyses of different molecular properties of the mitochondrial genomes showed an intricate network of direct and indirect correlations between the different molecular factors. Hence, gene order evolution seems to be driven by molecular evolutionary aspects rather than by life history or ecology. On the other hand, variability of the gene order does not predict if a taxon is difficult to place in molecular phylogenetic reconstructions using sequence data or not. We also discuss the molecular properties of annelid mitochondrial genomes considering canonical views on gene evolution and potential reasons why the canonical views do not always fit to the observed patterns without making some adjustments.
Background: The phylogenetic bootstrap, first proposed by Felsenstein in 1985, is a critically important statistical method in assessing the robusticity of phylogenetic datasets. Core to its concept was the use of pseudo sampling - assessing the data by generating new replicates derived from the initial dataset that was used to generate the phylogeny. In this way, phylogenetic support metrics could overcome the lack of perfect, infinite data. With infinite data, however, it is possible to sample smaller replicates directly from the data to obtain both the phylogeny and its statistical robusticity in the same analysis. Due to the growth of whole genome sequencing, the depth and breadth of our datasets have greatly expanded and are set to only expand further. With genome-scale datasets comprising thousands of genes, we can now obtain a proxy for infinite data. Accordingly, we can potentially abandon the notion of pseudo sampling and instead randomly sample small subsets of genes from the thousands of genes in our analyses. Methods: We introduce Scoutknife, a jackknife-style subsampling implementation that generates 100 datasets by randomly sampling a small number of genes from an initial large-gene dataset to jointly establish both a phylogenetic hypothesis and assess its robusticity. We assess its effectiveness by using 18 previously published datasets and 100 simulation studies. Results: We show that Scoutknife is conservative and informative as to conflicts and incongruence across the whole genome, without the need for subsampling based on traditional model selection criteria. Conclusions: Scoutknife reliably achieves comparable results to selecting the best genes on both real and simulation datasets, while being resistant to the potential biases caused by selecting for model fit. As the amount of genome data grows, it becomes an even more exciting option to assess the robusticity of phylogenetic hypotheses.
Fleming, James Frederick; Pisani, Davide & Arakawa, Kazuharu
(2023).
The evolution of temperature and desiccation-related protein families in Tardigrada reveals the complex acquisition of extremotolerance.
Genome Biology and Evolution (GBE).
16(1).
doi: 10.1093/gbe/evad217.
Full text in Research Archive
Fleming, James Frederick; Gracia, Alberto Valero & Struck, Torsten H
(2023).
Identifying and addressing methodological incongruence in phylogenomics: A review.
Evolutionary Applications.
16(6),
p. 1087–1104.
doi: 10.1111/eva.13565.
Full text in Research ArchiveShow summary
Abstract The availability of phylogenetic data has greatly expanded in recent years. As a result, a new era in phylogenetic analysis is dawning—one in which the methods we use to analyse and assess our data are the bottleneck to producing valuable phylogenetic hypotheses, rather than the need to acquire more data. This makes the ability to accurately appraise and evaluate new methods of phylogenetic analysis and phylogenetic artefact identification more important than ever. Incongruence in phylogenetic reconstructions based on different datasets may be due to two major sources: biological and methodological. Biological sources comprise processes like horizontal gene transfer, hybridization and incomplete lineage sorting, while methodological ones contain falsely assigned data or violations of the assumptions of the underlying model. While the former provides interesting insights into the evolutionary history of the investigated groups, the latter should be avoided or minimized as best as possible. However, errors introduced by methodology must first be excluded or minimized to be able to conclude that biological sources are the cause. Fortunately, a variety of useful tools exist to help detect such misassignments and model violations and to apply ameliorating measurements. Still, the number of methods and their theoretical underpinning can be overwhelming and opaque. Here, we present a practical and comprehensive review of recent developments in techniques to detect artefacts arising from model violations and poorly assigned data. The advantages and disadvantages of the different methods to detect such misleading signals in phylogenetic reconstructions are also discussed. As there is no one‐size‐fits‐all solution, this review can serve as a guide in choosing the most appropriate detection methods depending on both the actual dataset and the computational power available to the researcher. Ultimately, this informed selection will have a positive impact on the broader field, allowing us to better understand the evolutionary history of the group of interest.
Struck, Torsten H & Cerca, José
(2022).
Connecting micro- and macro-evolutionary research – extant cryptic species as systems to understand macro-evolutionary stasis.
In Monro, Alexandre K. & Mayo, Simon J. (Ed.),
Cryptic species - Morphological Stasis, Circumscription, and Hidden Diversity.
Cambridge University Press.
ISSN 9781316513644.p. 143–168.
doi: 10.1017/9781009070553.006.
Full text in Research ArchiveShow summary
‘Cryptic species’ are nowadays commonplace in systematics, and, with the application of barcoding approaches, increasing numbers of species are shown to be supposedly cryptic. However, the use of the term is inconsistent in the literature. Moreover, processes associated with the formation of cryptic species are generally not taken into account. These processes comprise (I) recent divergence, (II) parallel or (III) convergent phenotypic evolution, or (IV) phenotypic stasis. This causes confusion and imprecision in attempting to resolve the origins, evolutionary trajectories and ecological importance of cryptic species.
Here, we will briefly present these four processes based on a recently proposed conceptual framework for the definition of cryptic species. The four categories differ from each other in their temporal scale, phylogenetic context, and underlying selection. In focusing on the phenomenon of stasis, that is the absence of change over long geological periods, we discuss the overlap between stasis seen in the paleontological record and as observed in cryptic species, and we further consider that the challenge of integrating stasis into current evolutionary thinking is due to several different factors. These include microevolutionary research findings in extant species that seem to indicate abundant variation in both genetic markers and quantitative traits and development. Hence, there are contradictions between micro- and macroevolutionary scales. Cryptic species can provide study systems of extant species, which make it possible to bridge the gap between micro- and macroevolutionary research. Here we discuss this potential and provide an outlook on future research directions exploiting cryptic species.
Fleming, James Frederick & Struck, Torsten H
(2022).
nRCFV: A new, dataset-size-independent metric to quantify compositional heterogeneity in nucleotide and amino acid datasets.
Research Square.
ISSN 2693-5015.
doi: 10.21203/rs.3.rs-2354937/v1.
Full text in Research Archive
Ecdysozoans (Phyla Arthropoda, Kinorhyncha, Loricifera, Nematoda, Nematomorpha, Onychophora, Priapulida, Tardigrada) are invertebrates bearing a tough, periodically moulted cuticle that predisposes them to exceptional preservation. Ecdysozoans dominate the oldest exceptionally preserved bilaterian animal biotas in the early to mid-Cambrian (c. 520–508 Ma), with possible trace fossils in the latest Ediacaran (<556 Ma). The fossil record of Ecdysozoa is among the best understood of major animal clades and is believed to document their origins and evolutionary history well. Strikingly, however, molecular clock analyses have implied a considerably deeper Precambrian origin for Ecdysozoa, much older than their earliest fossils. Here, using an improved set of fossil calibrations, we performed Bayesian analyses to estimate an evolutionary time-tree for Ecdysozoa, sampling all eight phyla for the first time. Our results recover Scalidophora as the sister group to Nematoida + Panarthropoda (= Cryptovermes nov.) and suggest that the Ediacaran divergence of Ecdysozoa occurred at least 23 myr before the first potential ecdysozoan trace fossils. This finding is impervious to the use of all plausible phylogenies, fossil prior distributions, evolutionary rate models and matrix partitioning strategies. Arthropods exhibit more precision and less incongruence between fossil- and clock-based estimates of clade ages than other ecdysozoan phyla.
Open lectures on biodiversity from ants to ocean life, and how taxonomy helps us understand ecology and evolution. With Corrie S. Moreau, Cornell University, and Torsten H. Struck, University of Oslo.
Fleming, James Frederick & Struck, Torsten H
(2024).
BOSTIn – User friendly software to assess potential problems in genomic datasets.
Full text in Research Archive
Struck, Torsten H
(2024).
Species selection for large-scale genome projects – an automated process based on explicityly bottom-up defined criteria.
Full text in Research Archive
Eriksen, Pia Merete; Fleming, James Frederick & Struck, Torsten H
(2024).
Another step up the phylogenetic hill: Assessing the viability of macrosynteny as a phylogenetic trait in Lophotrochozoa.
Full text in Research Archive
Eriksen, Pia Merete; Fleming, James Frederick & Struck, Torsten H
(2024).
Evaluating the efficacy of macrosynteny as a phylogenetic trait in Lophotrochozoa.
Full text in Research Archive
Le, Yen Nhu Dinh; Fleming, James Frederick & Struck, Torsten H
(2024).
Partial polyploidy and potential heteroplasmy were revealed through the attempt at the whole mitochondrial genome study of Scoloplos cf. armiger.
Full text in Research Archive
Bachmann, Lutz & Struck, Torsten H
(2024).
The 2024 Master, PhD and PostDoc Days at the Natural History Museum Oslo – Blogs of the CEG research group.
Full text in Research Archive
Eriksen, Pia Merete & Struck, Torsten H
(2024).
Door 23: Struggles, Setbacks & Solutions – Blogs of the CEG research group.
Full text in Research Archive
Fleming, James Frederick & Melteig, Elina
(2024).
Blekkspruten er i slekt med snegler, men ingen vet nøyaktig hvordan.
[Internet].
https://www.titan.uio.no/naturvitenskap/2023/blekkspruten-er.
Full text in Research Archive
McGrath, Casey & Fleming, James Frederick
(2024).
Highlight: Tardigrades and the Science of Extreme Survival.
[Journal].
Genome Biology and Evolution (GBE).
doi: 10.1093/gbe/evad234.
Full text in Research Archive
Melteig, Elina & Fleming, James Frederick
(2024).
Entusiaster bruker ferien på å lete etter et dyr som kanskje ikke finnes.
[Journal].
https://www.titan.uio.no/naturvitenskap/2024/bruker-ferien-p.
Full text in Research Archive
Fry, Hannah; Briain, Dara Ó & Fleming, James Frederick
(2024).
Curious Cases Season 1 Episode 7. Invincible Moss Piglets.
[Radio].
https://www.bbc.co.uk/programmes/m002404x.
Full text in Research Archive
Fleming, James Frederick
(2024).
Group of the month: Loricifera.
[Internet].
https://blog.annelida.de/2024/04/15/group-of-the-month-loric.
Full text in Research Archive
Fleming, James Frederick
(2024).
Stories from Slime Eels: How the Hagfish Helps Us Understand Humans.
[Internet].
https://blog.annelida.de/2024/01/15/stories-from-slime-eels-.
Full text in Research Archive
Struck, Torsten H
(2024).
Tror de kan gjenopplive mammuten i 2028: – Nesten motsatt av «Jurassic Park».
[Journal].
NRK.
Full text in Research Archive
Gracia, Alberto Valero
(2023).
HiFi de novo sequencing and genome architecture dynamics in nemerteans.
Full text in Research Archive
Struck, Torsten H
(2023).
Multidimensional Phylogenetic Hypothesis Evaluation – a suggestion.
Full text in Research Archive
Struck, Torsten H; Helsem, Stian Aleksander; Dimitrov, Dimitar & Halanych, Kenneth M.
(2023).
Dating the annelid tree of life.
Full text in Research Archive
Nickellaus, Robert; Struck, Torsten H & Kocot, Kevin M.
(2023).
Whole Genome Amplification for de novo Genomics – the Genome of the Gastrotrich, Lepidodermella squamata as a case study.
Full text in Research Archive
Struck, Torsten H
(2023).
Species selection for genome projects such as ERGA and BGE – an semi-automated, bottom-up process based on explicitly defined criteria.
Full text in Research Archive
Gracia, Alberto Valero
(2023).
Group of the month: Rotifera – Blogs of the CEG research group.
Full text in Research Archive
RCFV reader is a new software to quickly calculate normalised relative compositional frequency variability. - JFFleming/RCFV_Reader
Capucho, Ana Teresa & Struck, Torsten H
(2023).
Group of the month: Nemertea – Blogs of the CEG research group.
Full text in Research Archive
Struck, Torsten H
(2023).
The chosen ones or how to select species to genome sequence – Blogs of the CEG research group.
Full text in Research Archive
Struck, Torsten H
(2023).
Doing genomic research with a masonry trowel – Blogs of the CEG research group.
Full text in Research Archive
Struck, Torsten H
(2023).
Group of the month: Parergodrilidae (Annelida) – Blogs of the CEG research group.
Full text in Research Archive
Struck, Torsten H
(2023).
ERGA pilot project – what can we learn for future genome projects – Blogs of the CEG research group.
Full text in Research Archive
Struck, Torsten H
(2023).
ERGA pilot project – the selected species – Blogs of the CEG research group.
Full text in Research Archive
Struck, Torsten H
(2023).
Systematics and ecosystem function – can they go together? – Blogs of the CEG research group.
Full text in Research Archive
Struck, Torsten H
(2023).
The mitochondrial gene order of Annelida – extremely old, but still working – Blogs of the CEG research group.
Full text in Research Archive
Struck, Torsten H
(2023).
The importance of the small stuff for science – Blogs of the CEG research group.
Full text in Research Archive
Fleming, James Frederick
(2023).
Tardigrades, Public Databases and Science From Your Settee.
[Internet].
https://blog.annelida.de/2023/03/07/tardigrades-public-datab.
Full text in Research Archive
Fleming, James Frederick
(2023).
The measure of our reach: understanding evolution when our models break down.
[Internet].
https://blog.annelida.de/2023/04/19/the-measure-of-our-reach.
Full text in Research Archive
Melteig, Elina & Fleming, James Frederick
(2023).
Dyret som gjenoppstår fra de døde – helt naturlig.
[Internet].
https://www.titan.uio.no/naturvitenskap/2023/dyret-som-gjeno.
Full text in Research Archive
Fleming, James Frederick
(2023).
On Pre-Prints, Publishing and Peer Review.
[Internet].
https://blog.annelida.de/2023/08/08/on-pre-prints-publishing.
Full text in Research Archive
Fleming, James Frederick
(2023).
Group of the Month: Placozoa.
[Internet].
https://blog.annelida.de/2023/07/01/group-of-the-month-placo.
Full text in Research Archive
Fleming, James Frederick
(2023).
Writing a Review: Introducing Your Field.
[Internet].
https://blog.annelida.de/2023/06/28/writing-a-review-introdu.
Full text in Research Archive
Fleming, James Frederick
(2023).
Scoutknife: Understanding Likelihood in an Era of Perfect Data.
Full text in Research Archive
Fleming, James Frederick
(2023).
Day 15: What’s coming next year!
[Internet].
https://blog.annelida.de/2023/12/15/day-15-whats-coming-next.
Full text in Research Archive
Fleming, James Frederick
(2023).
Day 14: When is a Larva like a Birthday Present?
[Internet].
https://blog.annelida.de/2023/12/14/day-14-when-is-a-larva-l.
Full text in Research Archive
Fleming, James Frederick
(2023).
Day 8: Working in Phylogenetic Methods.
[Internet].
https://blog.annelida.de/2023/12/08/day-8-working-in-phyloge.
Full text in Research Archive
The genomics revolution completely altered our understanding of phylogeny - the study of the relationships between animals. The identification of new cryptic species through phylogenetic methods that combine molecular and morphological data has led to an appreciation of phylogenetic relatedness as a biodiversity concept. Currently, however, phylogenetics has adopted a conservative stance towards this deluge of “big data”, selecting genes that fit well within our models of evolution - this has the benefit of evading artifactual topologies that violate these models, but may well be presenting us with hypotheses of evolution that are preselected according to our own biases, or incorrectly causing us to adopt great confidence in hypotheses that are not as well supported by the data as it first appears. This can have a powerful impact on our understanding of the relationships between taxa, which in turn could erroneously inform future sampling or conservation efforts.
From this perspective, new phylogenetic methods that are optimized to make the most of “big data” may be of critical importance to advancing the field. As part of the University of Oslo’s Invertomics project, a small part of the broader European effort to sequence every Eukaryotic genome on the continent, we have developed new methods to assess phylogenetic data and new software to reconstruct phylogenetics optimized for whole genome data. We are additionally in the process of applying that to real genomic challenges across the second largest invertebrate superphylum, Lophotrochozoa. This talk will cover three new software packages, one of which has already been released, that hope to advance our understanding of organismal relationships in this new era of data. It will also explore the challenges of applying big data approaches to organismal phylogenetics.
Fleming, James Frederick
(2023).
Scoutknife: Understanding Likelihood in an Era of Perfect Data.
Full text in Research Archive
Struck, Torsten H
(2022).
Abiotic conditions of sediment & a little bit about distribution.
Full text in Research Archive
Gracia, Alberto Valero
(2022).
Gnathostomulida, an obscure and often neglected phylum – Blogs of the CEG research group.
Full text in Research Archive
Fleming, James Frederick & Struck, Torsten H
(2022).
Group of the month: Tardigrades – Blogs of the CEG research group.
Full text in Research Archive
Struck, Torsten H
(2022).
Group of the month: hairybellies (Gastrotricha) – Blogs of the CEG research group.
Full text in Research Archive
Struck, Torsten H
(2022).
Launch of Biodiversity Genomics Europe – sequencing the genomes of European species – Blogs of the CEG research group.
Full text in Research Archive
Struck, Torsten H
(2022).
Door 17: The carbon footprint of our computational analyses – Blogs of the CEG research group.
Full text in Research Archive
Fleming, James Frederick & Struck, Torsten H
(2022).
nRCFV: a sequence, taxon and character state-normalised metric for the pre-reconstruction evaluation of compositional heterogeneity.
Full text in Research Archive
Fleming, James Frederick & Struck, Torsten H
(2021).
Science From Lockdown: Advent Calendar December 4th 2021 – Blogs of the CEG research group.
Full text in Research Archive
This work explores macrosynteny, that is, conservation of gene order on a chromosomal scale, and evaluates the boundaries in which it can be applied as phylogenetic trait, using superphylum Spiralia as a case study. Spiralia is a diverse group of invertebrates, spanning the familiar, such as earthworms and mussels, to the unfamiliar, such as the enigmatic arrow-worms (Chaetognatha) and hairybellies (Gastrotricha). This work stress-tests macrosynteny by applying it to this group, and testing the parameters under which it operates. These parameters include the human factors of annotation, and choice of analysis composition, to the biological, such as genome rearrangement. This work finds that macrosynteny can indeed return informative results, however it is best applied with nuance and an understanding of the genomic landscape of the taxa under evaluation.
Annelida is recognized as one of the most morphologically and ecologically diverse phyla within Lophotrochozoa. Besides Arthropoda and Chordata, it is one of three taxa traditionally considered as being segmented. With over 22,000 species identified across terrestrial, freshwater, and marine environments, annelids are ubiquitous organisms inhabiting almost every habitat on earth. Despite their dominance in terms of population and species diversity, annelids remain mysterious, particularly in genomic study, where many aspects of their genetic makeup are yet to be thoroughly explored. Although employing advanced sequencing technologies such as whole genomic sequencing has become popular, genomes are not available for many families of Annelida leading to the limited knowledge about the genomic evolution in Annelida. In this study, the genomic data for four annelid taxa Scoloplos cf. armiger, Trilobodrilus axi, Protodriloides chaetifer, and Protodriloides symbioticus was generated. To explore the genomic characteristics, I aimed to get high quality DNA for PacBio Revio sequencing by using different commercial kits such as DNeasy Tissue Kit, MagAttract HMW DNA Kit, Nanobind PanDNA Kit, and REPLI-g advanced DNA single cell kit. Due to lack of reference genomic data, the de no assembly was used to build up the entire genome of these species. However, the genome of Trilobodrilus axi, Protodriloides chaetifer, and Protodriloides symbioticus essentially failed so more research needs to be carried out to get retrieve a high-quality genome for them. Despite challenges, a high-quality nuclear genome and a potential heteroplasmic mitochondrial genome were characterized in Scoloplos cf. armiger. The mitochondrial genome had a highly variable region between trnC and trnL2 across the different variants found in the specimen. These finding can contribute a valuable genomic resource for Annelida, enabling future comparative studies and phylogenetic reconstructions.